Data Portals
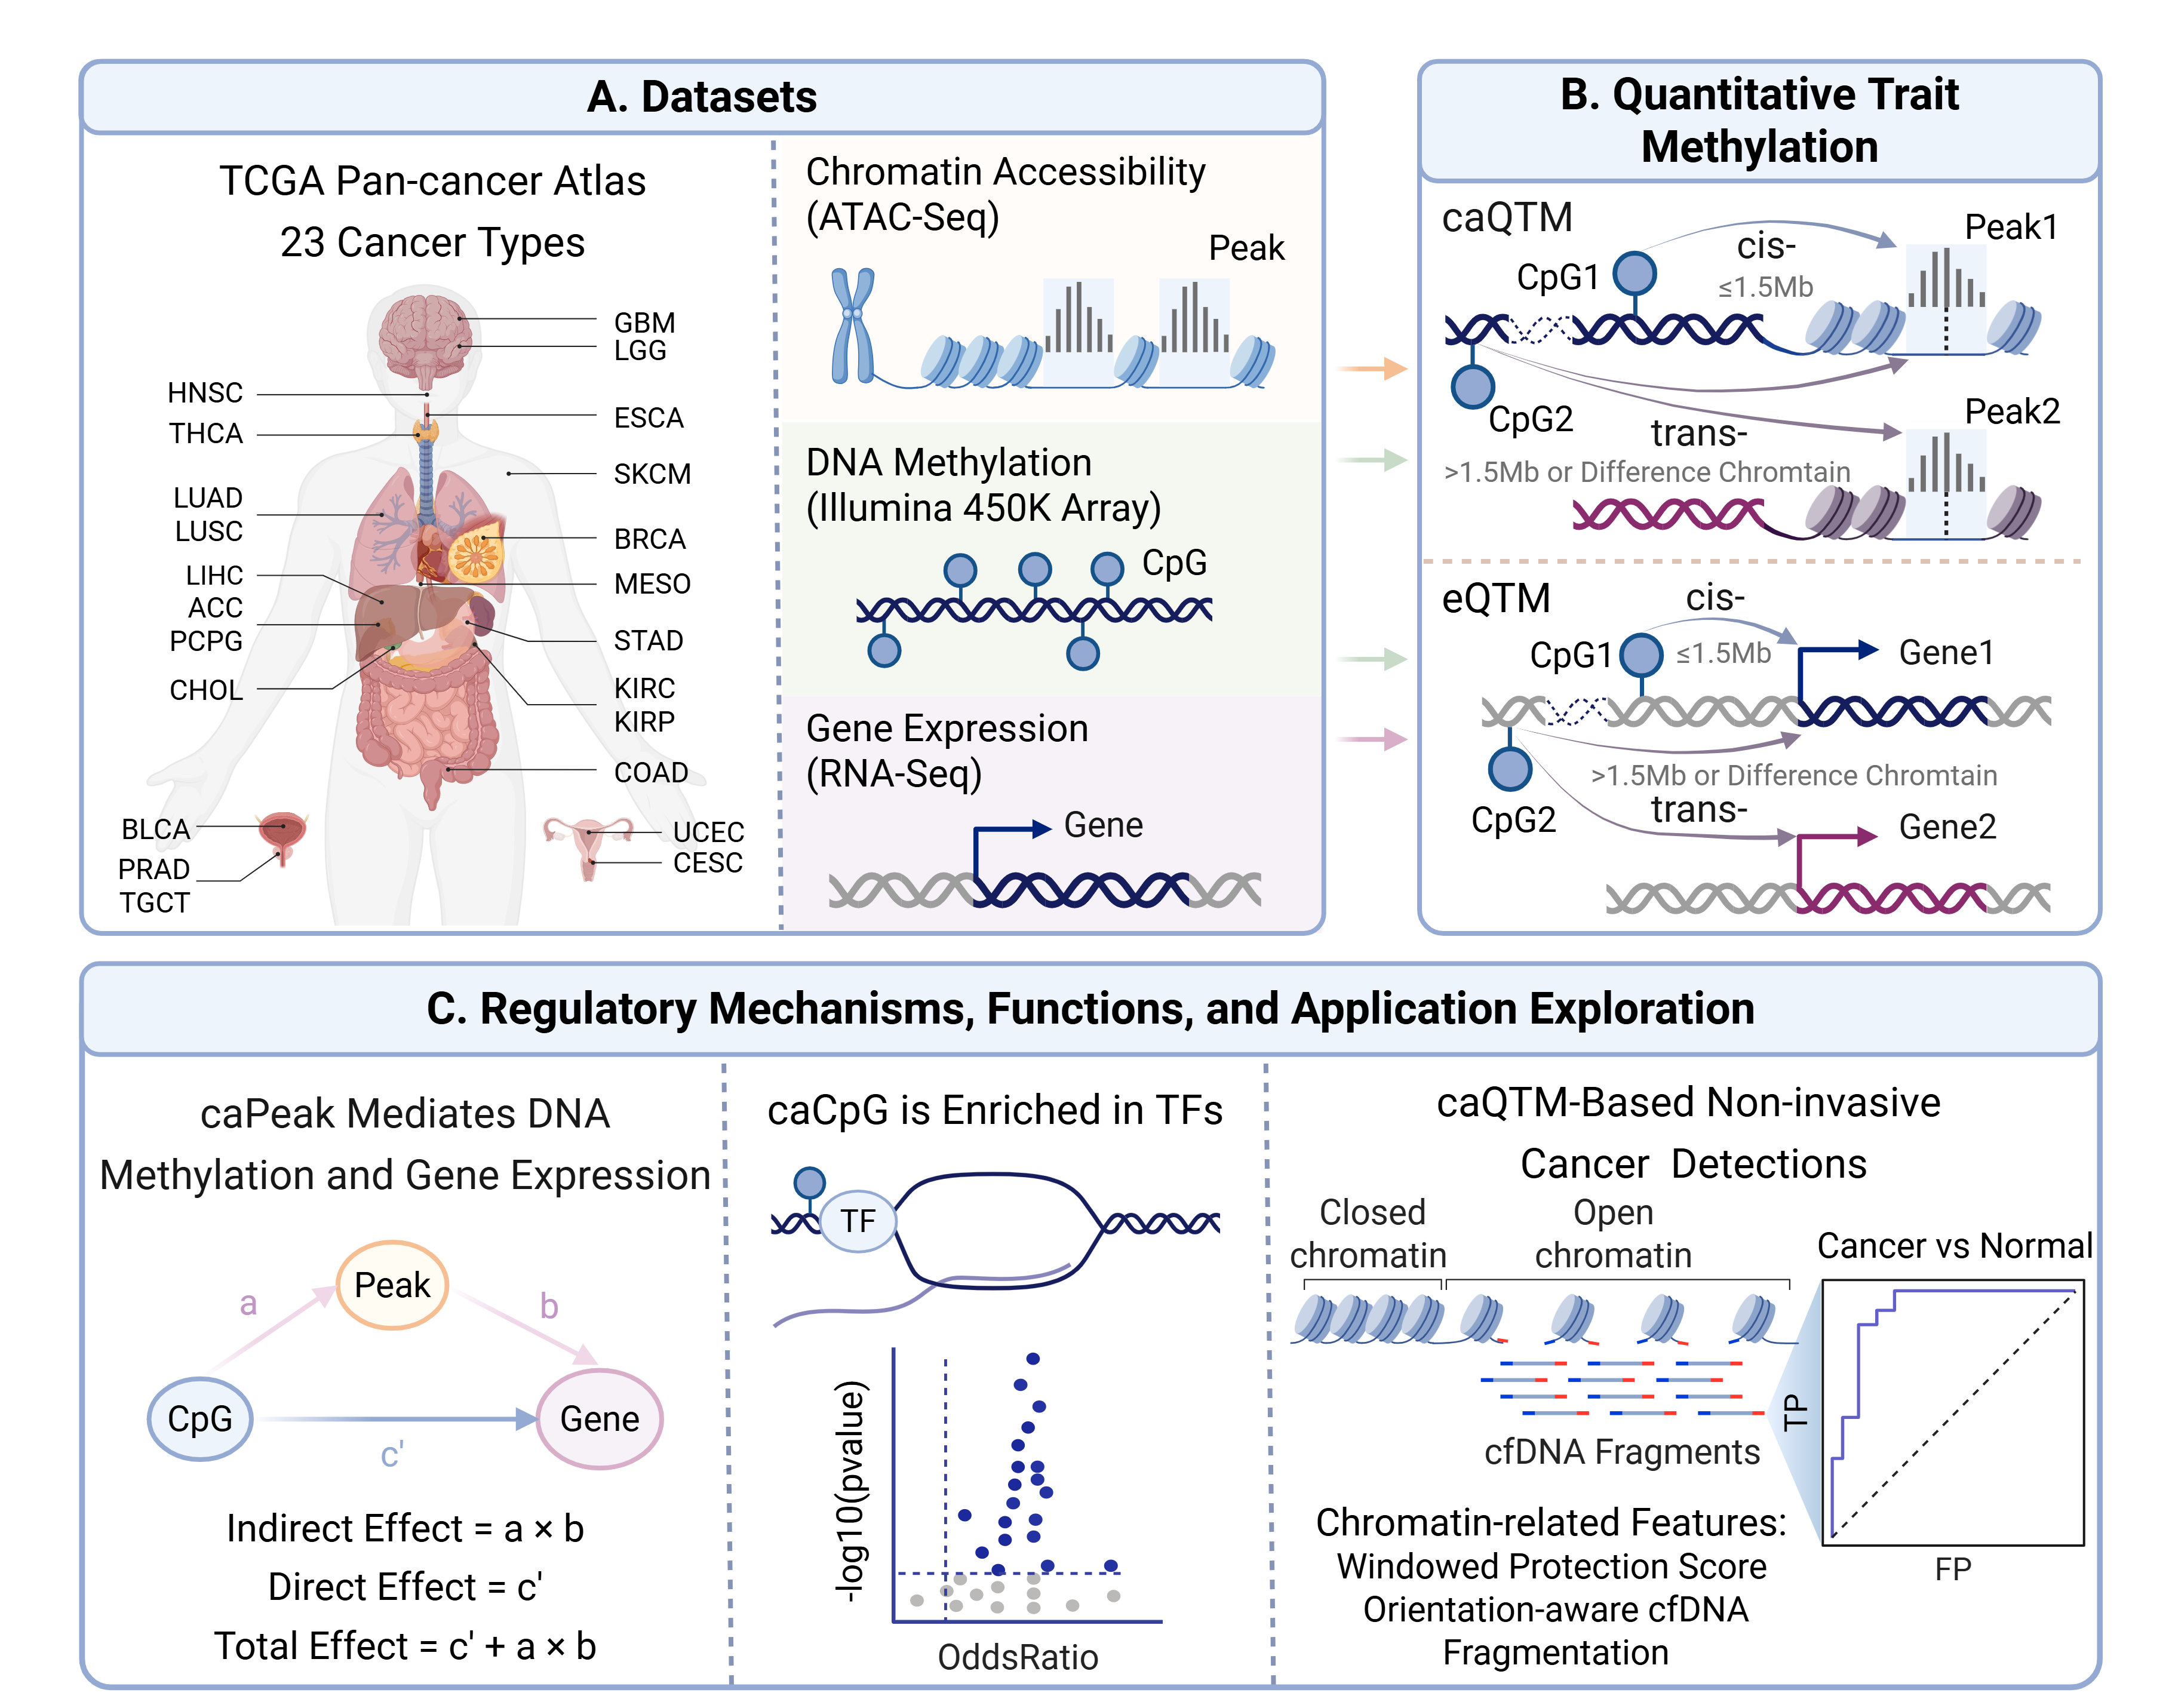
Pan-cancer caQTM: Pan-cancer association study of DNA methylation and chromatin accessibility
DNA methylation and chromatin accessibility are key epigenetic mechanisms regulating gene transcription, yet their coordinated roles in gene regulation remain poorly understood. We integrated multi-omics data from 23 TCGA cancer types to construct the first pan-cancer chromatin accessibility quantitative trait methylation (caQTM) atlas, identifying 157,596 significant methylation–accessibility associations. By combining caQTM, expression quantitative trait methylation (eQTM), and mediation analyses, we identified 5,457 significant CpG–caPeak–eGene regulatory chains, supporting a potential mediating role of chromatin accessibility in methylation-associated transcriptional regulation. Transcription factor enrichment analyses further suggested that caQTMs are involved in transcription factor–mediated chromatin remodeling. Finally, leveraging cancer-associated epigenetic loci, we demonstrated that functional caPeak regions can serve as informative cfDNA fragmentomic features, achieving an AUC of 0.88 for both breast and lung cancer classification. Together, our study establishes a pan-cancer caQTM landscape, reveals widespread links among DNA methylation, chromatin accessibility, and gene expression, and highlights functionally annotated regulatory regions as promising candidates for cfDNA-based cancer biomarkers.
Explore Database →